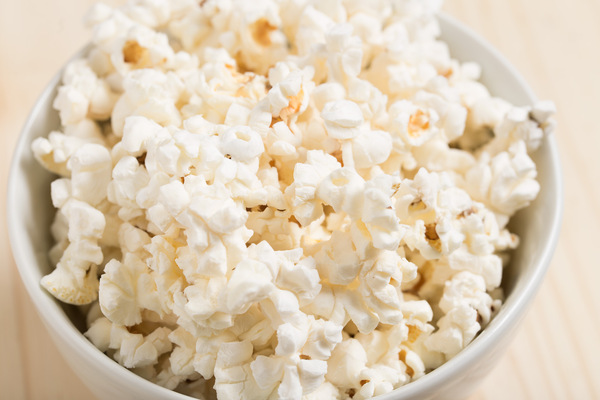
Have you ever wondered what contributes to the popping ability of popcorn? In this study, the authors use Quantitative Trait Locus (QTL) mapping to identify genes that may contribute to specific popping characteristics including kernel size and popping expansion volume (PEV).
The authors assess a genetic variant within a well-known interaction partner of huntingtin that has been linked to modifying the age of onset of Huntington's disease.
Here, seeking to better understand the genetic associations underlying non-small cell lung cancer, the authors screened hundreds of genes, identifying that KCNMB2 upregulation was significantly correlated with poor prognoses in lung cancer patients. Based on this, they used small interfering RNA to decrease the expression of KCNMB2 in A549 lung cancer cells, finding decreased cell proliferation and increased lung cancer cell death. They suggest this could lead to a new potential target for lung cancer therapies.
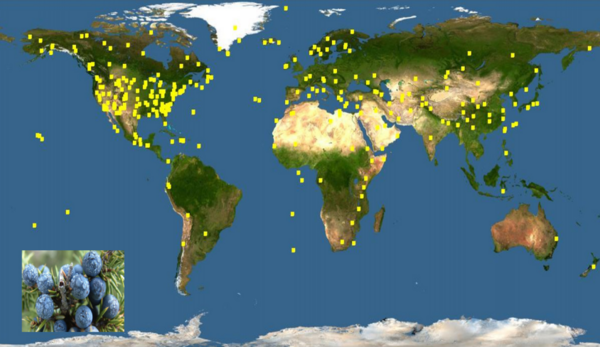
Many species of trees are distributed widely around the world, though not always in a way that makes immediate sense. The authors here use genetic information to help explain the geographic distribution of various conifer species throughout the world.
Although the United States maintains millions of square kilometers of nature reserves to protect the biodiversity of the specimens living there, little is known about how confining these species within designated protected lands influences the genetic variation required for a healthy population. In this study, the authors sequenced genetic barcodes of insects from a recently established nature reserve, the Southwestern Riverside County Multi-Species Reserve (SWRCMSR), and a non-protected area, the Mt. San Jacinto College (MSJC) Menifee campus, to compare the genetic variation between the two populations. Their results demonstrated that the midge fly population from the SWRCMSR had fewer unique DNA barcode sequence changes than the MSJC population, indicating that the comparatively younger nature reserve's population had likely not yet established its own unique genetic drift changes.
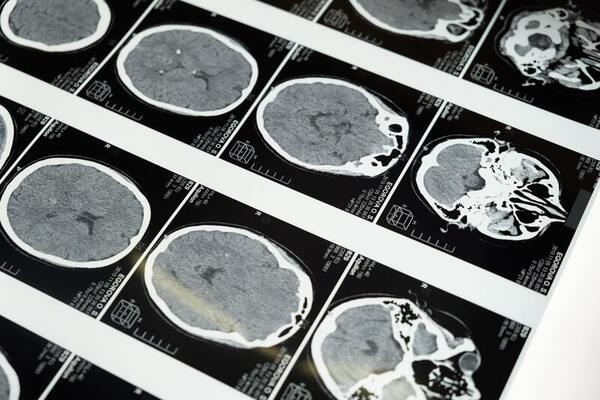
Glioblastoma Multiforme (GBM) is the most malignant brain tumor with the highest fraction of genome alterations (FGA), manifesting poor disease-free status (DFS) and overall survival (OS). We explored The Cancer Genome Atlas (TCGA) and cBioportal public dataset- Firehose legacy GBM to study DNA repair genes Activating Signal Cointegrator 1 Complex Subunit 3 (ASCC3) and Alpha-Ketoglutarate-Dependent Dioxygenase AlkB Homolog 3 (ALKBH3). To test our hypothesis that these genes have correlations with FGA and can better determine prognosis and survival, we sorted the dataset to arrive at 254 patients. Analyzing using RStudio, both ASCC3 and ALKBH3 demonstrated hypomethylation in 82.3% and 61.8% of patients, respectively. Interestingly, low mRNA expression was observed in both these genes. We further conducted correlation tests between both methylation and mRNA expression of these genes with FGA. ASCC3 was found to be negatively correlated, while ALKBH3 was found to be positively correlated, potentially indicating contrasting dysregulation of these two genes. Prognostic analysis showed the following: ASCC3 hypomethylation is significant with DFS and high ASCC3 mRNA expression to be significant with OS, demonstrating ASCC3’s potential as disease prediction marker.
In this study, the authors identify transcripts and gene networks that are changed after infection with the Middle East Respiratory Syndrome-related coronavirus (MERS-CoV).